Conditions · Metabolism: carbohydrates and glycogen
Andersen disease (glycogen storage disease type IV) !
In Andersen disease glycogen forms with long, abnormal branches that damage the liver and lead to cirrhosis in childhood.
Updated 2026-11-01
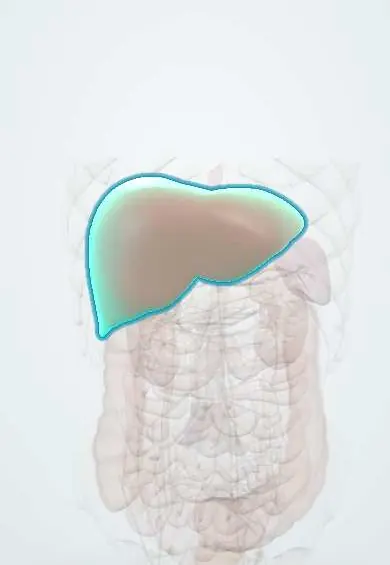
Structures involved
Tap one to see what it is and open it in 3D.
In biochemistry
What it is
Inherited glycogen storage disease caused by lack of the glycogen branching enzyme (GBE1 gene). A poorly branched glycogen (polyglucosan) builds up and the body treats it as foreign.
Who gets it
Very rare: 1 in 600,000 to 800,000 people, about 3 % of glycogen storage diseases.
Causes
- Autosomal recessive inheritance: GBE1 gene variants
Risk factors
- Carrier parents or affected siblings
Pathophysiology
Without the branching enzyme, glycogen synthase makes long chains with almost no branches. This polyglucosan is poorly soluble, deposits in hepatocytes and other cells, causes inflammation and fibrosis and leads to cirrhosis. The amount of glycogen is not much increased: the problem is its shape. In neuromuscular forms it deposits in muscle, heart and nerves.
Classification
- Classic progressive liver form: cirrhosis in the first years
- Non-progressive liver form
- Neuromuscular forms: severe perinatal, congenital, childhood or adult (polyglucosan body disease)
Symptoms and signs
- Failure to thrive in the first year
- Enlarged liver and spleen
- Ascites and varices from portal hypertension
- Weakness and hypotonia in the neuromuscular forms
Diagnosis
- Clinical
- Infant with failure to thrive, hepatosplenomegaly and signs of chronic liver disease.
- Tests
- Raised transaminases and abnormal liver function; hypoglycaemia appears late. Liver biopsy with polyglucosan deposits (PAS positive, diastase resistant). Low branching enzyme activity and GBE1 testing.
- Imaging
- Abdominal ultrasound with hepatosplenomegaly, signs of cirrhosis and portal hypertension.
Treatment
- Surgical
- Liver transplant in the progressive liver form
- Lifestyle
- Nutritional support and frequent meals to avoid hypoglycaemia
- Medical
- Management of cirrhosis complications
- Follow-up
- Monitoring of heart and neuromuscular function
Complications
- Cirrhosis and liver failure
- Portal hypertension with varices
- Cardiomyopathy
Prognosis
Without transplant, the classic liver form usually causes liver failure before age 5.
Test yourself
Think of the answer, then tap the question to see it.
Which enzyme is missing in Andersen disease?
The glycogen branching enzyme (GBE1).
What is abnormal about glycogen in this disease?
It has long chains and few branches (polyglucosan), is poorly soluble and damages cells.
What is the definitive treatment of the progressive liver form?
Liver transplant.
Related conditions
Educational information, not medical advice. For any symptom, see a health professional.
Sources: Glycogen storage disease type IV: MedlinePlus Genetics (MedlinePlus, US National Library of Medicine, 2026); GBE1 gene: MedlinePlus Genetics (MedlinePlus, US National Library of Medicine, 2026); Textbook of Biochemistry for Medical Students, 6th ed. (Jaypee Brothers Medical Publishers (Vasudevan DM, Sreekumari S, Vaidyanathan K), 2011).
Spotted a mistake?We are always improving for the community. Tell us what is wrong and our team will review it.Report a mistake
Representative model, for educational purposes only.