Conditions · Metabolism (biochemistry)
Pyruvate dehydrogenase deficiency !
Pyruvate dehydrogenase deficiency is a genetic disease in which cells cannot burn sugar properly; it mainly harms the brain.
Updated 2026-10-31
Structures involved
Tap one to see what it is and open it in 3D.
In biochemistry
- Krebs cycle: Link (PDH) · Pyruvate dehydrogenase complex (PDH)
- Alanine
- Acetyl-CoA
- Pyruvate dehydrogenase complex (PDH)
What it is
Low activity of the pyruvate dehydrogenase complex, usually from PDHA1 variants on the X chromosome. Pyruvate cannot enter the Krebs cycle and builds up as lactate.
Who gets it
It is one of the most common causes of congenital lactic acidosis; it affects boys and girls, with more variable pictures in girls because of X inactivation.
Causes
- PDHA1 variants (X-linked), most of them new
- Less often: PDHB, DLAT, DLD, PDHX or the PDP1 phosphatase (autosomal recessive)
Risk factors
- Affected siblings or a carrier mother
Pathophysiology
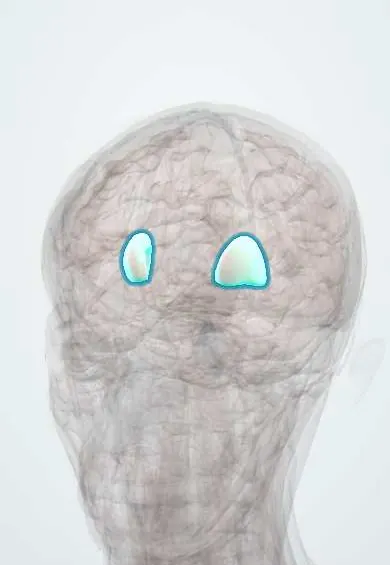
Without PDH the brain, which depends on glucose, gets too little acetyl-CoA and ATP. Pyruvate is diverted to lactate and alanine. The basal ganglia and brainstem are damaged (Leigh syndrome) and brain development may be affected before birth. The lactate/pyruvate ratio stays normal because the respiratory chain works.
Classification
- Neonatal form: severe lactic acidosis
- Infantile form: developmental delay, Leigh syndrome
- Mild form: episodic ataxia after carbohydrate-rich meals
Symptoms and signs
- Fast breathing, lethargy and poor feeding in the newborn
- Low muscle tone and developmental delay
- Seizures
- Ataxia that worsens with fever or carbohydrates
- Abnormal eye movements
Diagnosis
- Clinical
- Lactic acidosis with neurological signs, especially if it worsens with glucose.
- Tests
- Blood and spinal fluid lactate and pyruvate (normal ratio, under 20), high alanine; PDH activity in fibroblasts or lymphocytes; gene panel (PDHA1 and others).
- Imaging
- Brain MRI: symmetric basal ganglia and brainstem lesions (Leigh), absent corpus callosum or cysts; spectroscopy shows a lactate peak.
Treatment
- Lifestyle
- Ketogenic diet: supplies ketones as fuel that bypasses PDH
- Medical
- Thiamine, because some variants respond to it
- Medical
- Dichloroacetate in selected cases (inhibits PDH kinase)
- Acute
- In crises: correct acidosis and avoid large glucose loads
Complications
- Intellectual disability
- Epilepsy
- Respiratory failure during crises
Test yourself
Think of the answer, then tap the question to see it.
How is the lactate/pyruvate ratio in PDH deficiency?
Normal (under 20), unlike respiratory-chain defects.
Why is the ketogenic diet used?
Ketones enter the cycle as acetyl-CoA without going through PDH.
Which gene causes most cases and how is it inherited?
PDHA1, X-linked.
Related conditions
Educational information, not medical advice. For any symptom, see a health professional.
Sources: Pyruvate dehydrogenase deficiency: MedlinePlus Genetics (MedlinePlus, US National Library of Medicine, 2026); PDHA1 gene: MedlinePlus Genetics (MedlinePlus, US National Library of Medicine, 2026); Leigh syndrome: MedlinePlus Genetics (MedlinePlus, US National Library of Medicine, 2026).
Spotted a mistake?We are always improving for the community. Tell us what is wrong and our team will review it.Report a mistake
Representative model, for educational purposes only.